The FDA grants Priority Review to LEO Pharma’s dersimelagon NDA for EPP and XLP, with a PDUFA action date set for February 2027.
Written By: Umesh Hanumante,
M.Pharm (Reg. Affairs)
Reviewed By: Pharmacally Editorial Team
The FDA has accepted LEO Pharma’s New Drug Application (NDA) for dersimelagon for filing and granted Priority Review for the investigational oral MC1R agonist in erythropoietic protoporphyria (EPP) and X-linked protoporphyria (XLP). The regulatory milestone comes as LEO Pharma confirms the closing of its acquisition of worldwide rights to the asset from Tanabe Pharma.
FDA Sets February 2027 PDUFA Date
The U.S. Food and Drug Administration has assigned a Prescription Drug User Fee Act (PDUFA) target action date by the end of February 2027. Priority Review establishes a six-month review goal for eligible applications, compared with the standard review goal of 10 months.
Dersimelagon has also previously received Fast Track Designation and Orphan Drug Designation from the FDA.
The therapy remains investigational, and its safety and efficacy have not been established by any regulatory authority.
Oral MC1R Agonist Addresses Photosensitivity
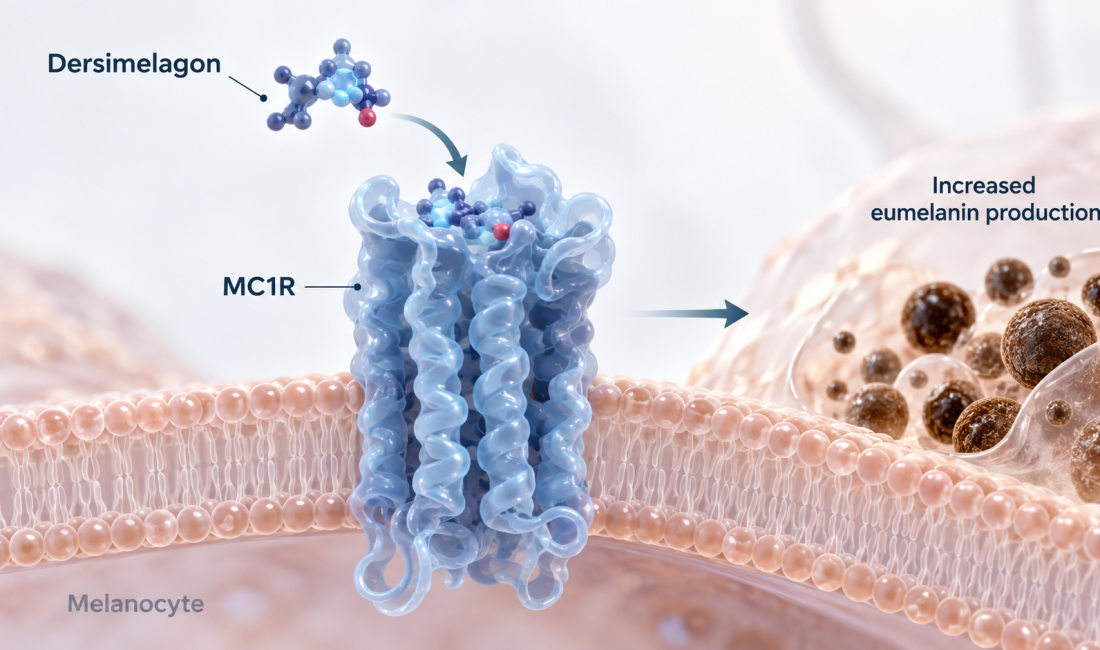
Dersimelagon is an investigational, once-daily oral small-molecule agonist of the melanocortin 1 receptor (MC1R). Activation of MC1R promotes melanogenesis and increases eumelanin production, potentially improving the skin’s tolerance to ultraviolet and visible light exposure.
EPP and XLP are rare inherited metabolic disorders caused by defects in heme biosynthesis that lead to protoporphyrin accumulation. Exposure to visible light can trigger severe phototoxic burning pain, often initially without obvious skin lesions. Patients may substantially restrict outdoor activities to avoid symptoms, while some can develop hepatic complications.
Management relies primarily on strict light avoidance and photoprotection, highlighting the need for additional treatment options that can improve patients’ ability to tolerate light exposure.
Phase 3 INSPIRE Trial Supports Filing
The NDA submission is supported by data from the global, randomized, double-blind, placebo-controlled Phase 3 INSPIRE study, presented as a late-breaking abstract at the 2026 AAD Annual Meeting.
Dersimelagon demonstrated statistically significant and clinically meaningful outcomes across the study’s primary and secondary endpoints. A key functional outcome was a significant prolongation in average daily sunlight exposure time before patients experienced their first prodromal symptoms.
Clinical updates have reported that dersimelagon was generally well tolerated. Reported adverse events included benign skin nevi, headache, nausea, diarrhea, and skin hyperpigmentation.
LEO Pharma Completes Worldwide Rights Acquisition
Concurrently, LEO Pharma completed its acquisition of worldwide rights to dersimelagon from Tanabe Pharma after fulfilling customary closing conditions.
The transaction structure, originally announced on August 18, 2026, remains unchanged. It provides for up to $435 million in upfront and near-term milestone payments, plus potential downstream milestones and tiered royalties ranging from double-digit percentages to the mid-teens on net sales.
The acquisition strengthens LEO Pharma’s late-stage rare dermatology pipeline and expands its global dermatology portfolio.
February 2027 Regulatory Decision in Focus
The FDA’s PDUFA action date by the end of February 2027 represents the next major milestone for dersimelagon. If approved, the therapy could become the first oral treatment for patients with EPP and XLP, providing a potential systemic option for conditions associated with severe sunlight-induced pain and substantial restrictions on daily life.
The FDA review remains ongoing, and the final benefit-risk assessment will determine whether dersimelagon can advance to regulatory approval.
Reference
LEO Pharma announces FDA acceptance of dersimelagon NDA with Priority Review and closes acquisition from Tanabe Pharma, LEO Pharma, September 18 2026
About the Writer
Umesh Hanumante (M.Pharm) (LinkedIn) is a pharmacy professional and healthcare writer with a background in Regulatory Affairs, pharmaceutical innovation, and clinical research. He has around two years of industry experience as an Executive PMT at Troikaa Pharmaceuticals Ltd and qualified GPAT 2024. His areas of interest include regulatory compliance, dossier preparation, clinical trials, emerging therapies, and advancements in the global pharmaceutical and healthcare sector.