Cullinan Therapeutics initiates FDA NDA submission for zipalertinib plus chemotherapy in first-line EGFR exon 20 insertion-mutated NSCLC.
Written By: Mayuresh Salvi, PharmD
Reviewed By: Pharmacally Editorial Team
Cullinan Therapeutics announced on October 1, 2026, that it had initiated the NDA submission process to the U.S. Food and Drug Administration (FDA) for zipalertinib combined with platinum-based chemotherapy in patients with previously untreated, locally advanced or metastatic NSCLC harboring epidermal growth factor receptor (EGFR) exon 20 insertion (ex20ins) mutations.
The submission process was initiated following an agreement with the FDA to use the Real-Time Oncology Review (RTOR) program. Under RTOR, sponsors can provide clinical data to the FDA before the complete application is submitted, allowing the agency to begin reviewing efficacy and safety information earlier in the application process.
Cullinan expects to complete the NDA submission by the end of 2026. The current submission is for the first-line combination regimen and remains investigational. Zipalertinib has not been approved by any health authority.
Phase 3 REZILIENT3 Supports Regulatory Submission
The NDA submission is based on results from the Phase 3 REZILIENT3 trial (NCT05973773), which evaluated zipalertinib plus platinum-based chemotherapy against chemotherapy alone as first-line treatment for patients with EGFR exon 20 insertion-mutated NSCLC.
The global study enrolled 285 patients, including six patients in the safety lead-in and 279 patients subsequently randomized to the combination or chemotherapy-alone groups. The randomized portion included previously untreated patients with locally advanced or metastatic nonsquamous NSCLC harboring EGFR exon 20 insertion mutations.
REZILIENT3 met its primary endpoint of progression-free survival (PFS). At the planned interim efficacy analysis after 122 PFS events, median PFS was 14.5 months with zipalertinib plus chemotherapy compared with 8.5 months with chemotherapy alone, corresponding to a hazard ratio of 0.50 (95% CI, 0.34–0.73; P=0.00015). This represented a 6.0-month improvement in median PFS.
The objective response rate was also higher with the combination, at 65.0% compared with 40.3% with chemotherapy alone (P<0.0001). Median duration of response was 14.2 months versus 9.9 months, respectively.
At an interim overall survival analysis with approximately 30% event maturity, the hazard ratio for death was 0.72 (95% CI, 0.42–1.23). Overall survival follow-up remains ongoing.
The REZILIENT3 findings were presented during a Presidential Symposium at the International Association for the Study of Lung Cancer (IASLC) 2026 World Conference on Lung Cancer (WCLC). Cullinan reported that the combination demonstrated the longest median PFS observed to date in the first-line EGFR exon 20 insertion-mutated NSCLC setting.
Zipalertinib Development Program
Zipalertinib, also known by the development codes CLN-081/TAS6417, is an orally available small molecule designed to target activating mutations in EGFR. The molecule was selected for its ability to inhibit EGFR variants containing exon 20 insertion mutations.
Zipalertinib is designed as a next-generation, irreversible EGFR inhibitor for a genetically defined subset of patients with NSCLC. The drug is being developed by Taiho Oncology, Inc., its parent company Taiho Pharmaceutical Co., Ltd., and in collaboration with Cullinan Therapeutics in the United States.
Zipalertinib remains investigational for the proposed first-line combination indication and has not been approved by any health authority.
Separate FDA Review for Previously Treated NSCLC
The first-line combination NDA is separate from an earlier regulatory application for zipalertinib monotherapy in patients with locally advanced or metastatic EGFR exon 20 insertion-mutated NSCLC who have previously received platinum-based chemotherapy.
Zipalertinib has received FDA Breakthrough Therapy designation for this previously treated population. An NDA seeking accelerated approval for zipalertinib monotherapy remains under FDA review, with a Prescription Drug User Fee Act (PDUFA) target action date of February 27, 2027.
The two regulatory efforts therefore address different treatment settings. The earlier application concerns zipalertinib monotherapy after prior platinum-based chemotherapy, while the newly initiated RTOR submission concerns zipalertinib combined with platinum-based chemotherapy as first-line treatment for previously untreated patients.
EGFR Exon 20 Insertion-Mutated NSCLC
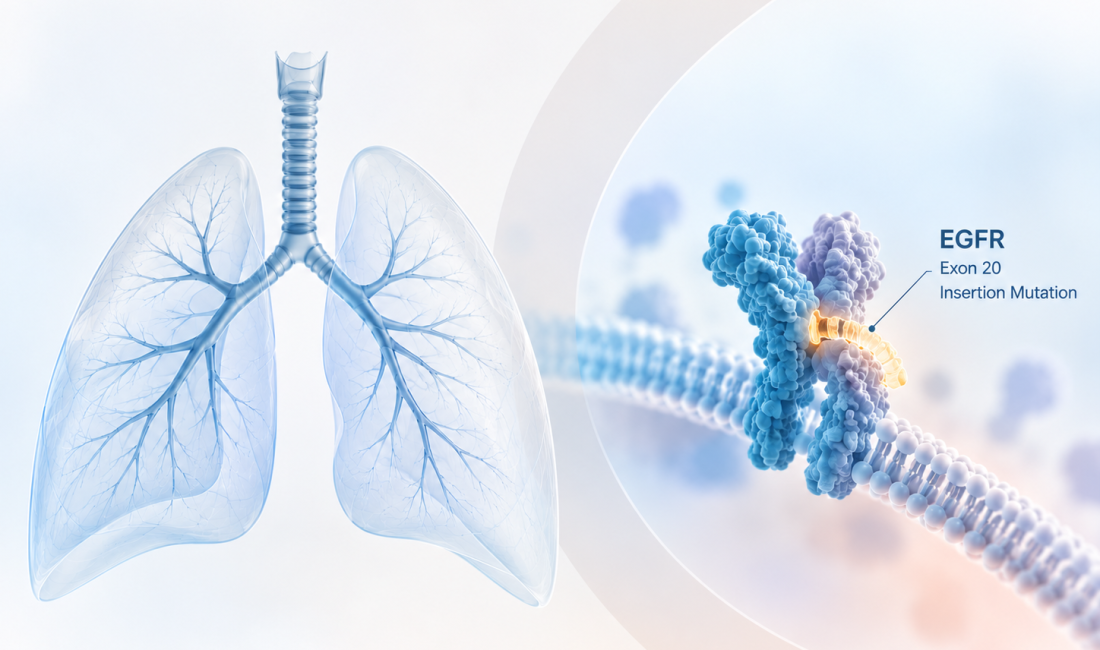
EGFR exon 20 insertion mutations represent a molecularly defined subset of NSCLC. Cullinan reports that EGFR exon 20 insertions occur in up to 4% of NSCLC cases globally. In the United States, approximately 16% of patients with NSCLC harbor EGFR mutations, with exon 20 insertions accounting for up to 12% of these EGFR mutations.
These estimates are supported by published epidemiological and genomic studies cited by Cullinan, including a systematic literature review and a genomic profiling study of EGFR exon 20 insertion mutations.
Regulatory Path Ahead
The initiation of the RTOR submission process represents the next regulatory step for zipalertinib plus platinum-based chemotherapy in previously untreated patients with locally advanced or metastatic EGFR exon 20 insertion-mutated NSCLC.
Cullinan expects to complete the NDA submission by the end of 2026. The FDA will subsequently determine whether the submitted clinical and regulatory evidence supports approval of the proposed first-line combination regimen.
Meanwhile, the separate NDA for zipalertinib monotherapy in previously treated EGFR exon 20 insertion-mutated NSCLC remains under FDA review, with a PDUFA target action date of February 27, 2027.
Reference
New Drug Application Submission Initiated for Zipalertinib Plus Chemotherapy in First-Line EGFR Exon 20 Insertion Mutation NSCLC for Review Under FDA Real-Time Oncology Review Program, Cullinan, 01 October 2026
REZILIENT3 (REsearching ZIpaLertinib In Egfr Non-small Cell Lung Cancer Tumors) (REZILIENT3), ClinicalTrials.gov ID NCT05973773
About the Writer
Mayuresh Sunil Salvi (Linkedin) is a PharmD professional and healthcare writer with a strong interest in pharmacovigilance, drug safety, and emerging medical research. He is passionate about exploring new drug discoveries, clinical research, and advances in evidence-based medicine. His interests also include ward rounds, prescription audits, and treatment analysis to support rational pharmacotherapy and improved patient care.