FDA approves a Boxed Warning for Adzynma over neutralizing ADAMTS13 antibodies linked to reduced treatment response and serious outcomes, including death.
Written By: Kirti Kumbhar, M. Pharm (QA)
Reviewed By: Pharmacally Editorial Team
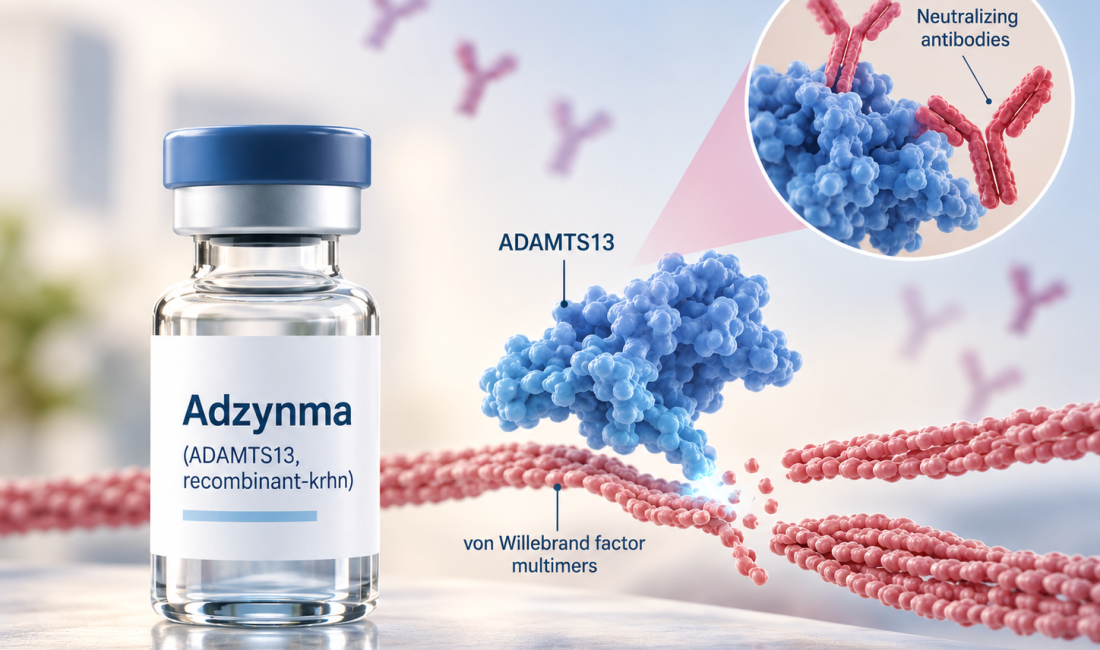
The U.S. Food and Drug Administration (FDA) has approved a revised label for Takeda Pharmaceuticals’ Adzynma (ADAMTS13, recombinant-krhn), adding a Boxed Warning for neutralizing antibodies against ADAMTS13 that can reduce or eliminate treatment response and have been associated with serious outcomes, including death. The September 23, 2026 action formally completes a regulatory process that began with an FDA safety investigation in November 2025.
From Safety Signal to Boxed Warning
The FDA’s action follows a stepwise safety and regulatory process:
- November 21, 2025: FDA reported postmarketing cases of neutralizing antibodies to ADAMTS13, including a fatal pediatric case, and said it was investigating the risk and evaluating whether further regulatory action was needed. At that time, the existing label already described the potential for neutralizing antibodies but did not include the postmarketing reports of serious or fatal outcomes.
- February 5, 2026: FDA formally notified Takeda that safety-related labeling changes were required under Section 505(o)(4) of the Federal Food, Drug, and Cosmetic Act. Takeda subsequently submitted a supplement proposing changes to the approved labeling.
- May 20, 2026: FDA issued a Safety Labeling Change Order requiring changes addressing neutralizing antibodies and their association with serious outcomes, including death. FDA and Takeda then worked through proposed labeling language.
- September 23, 2026: FDA approved Takeda’s supplemental BLA, including the Boxed Warning and updates to the Warnings and Precautions, Immunogenicity and Patient Information sections.
Neutralizing Antibodies Can Compromise Treatment
Adzynma is a recombinant ADAMTS13 enzyme replacement therapy approved for prophylactic or on-demand treatment of congenital thrombotic thrombocytopenic purpura (cTTP) in adults and pediatric patients. The rare inherited disorder results from severe ADAMTS13 deficiency, which can lead to uncontrolled platelet-rich microvascular thrombosis.
The concern is immunogenicity. Patients can develop antibodies that neutralize ADAMTS13 activity, potentially producing a decreased or absent response to recombinant or plasma-derived ADAMTS13. The revised FDA labeling states that neutralizing antibodies were not reported in the cTTP clinical trials, while postmarketing cases have now been associated with serious outcomes, including death.
Pediatric Death Triggered Regulatory Action
The original FDA safety communication described a 7-year-old patient who developed progressively worsening neurologic symptoms. Neutralizing antibodies to ADAMTS13 were identified approximately 10 months after prophylactic Adzynma treatment began. The patient had previously experienced severe allergic reactions to fresh frozen plasma.
FDA’s postmarketing surveillance identified the safety signal after Adzynma entered clinical use. The agency subsequently required Takeda to strengthen the product labeling and monitor the risk through additional postmarketing requirements.
Monitoring Added to the Boxed Warning
The newly approved labeling calls for close monitoring of all patients for ADAMTS13 activity and development of ADAMTS13 neutralizing antibodies. Testing is recommended before treatment begins and periodically during therapy.
Patients should not discontinue Adzynma without consulting their healthcare provider. The updated patient information also instructs patients to seek immediate medical attention for symptoms such as rash, hives, throat or chest tightness, difficulty breathing, dizziness, nausea or fainting.
FDA Finalizes the Revised Label
FDA approved the Package Insert and Patient Package Insert submitted by Takeda on September 17, 2026. The company must submit the final labeling in Structured Product Labeling format within 14 days of the September 23 approval letter.
The September action therefore represents the final labeling approval, rather than a newly discovered safety signal. FDA first disclosed the postmarketing risk in November 2025, formally required labeling changes in February and May 2026, and has now approved the final label carrying the Boxed Warning.
Reference
FDA Takes Action on New Boxed Warning for Adzynma Enzyme Replacement Therapy for Congenital Thrombotic Thrombocytopenic Purpura (cTTP), FDA. 23 September 2026
Approval Letter, 23 September 2026
About the Writer
Kirti Kumbhar (LinkedIn) is an M.Pharm graduate with experience in Quality Assurance at Lupin Limited and a strong interest in clinical research, regulatory affairs, and Trial Master File (TMF) management. She has developed knowledge of regulatory documentation, quality systems, compliance, and healthcare research through her professional experience. Passionate about clinical development and continuous learning, Kirti is committed to supporting high-quality healthcare documentation, regulatory excellence, and research-driven healthcare advancements.